This recall involves removing certain devices from where they are used or sold. The FDA has identified this recall as the most serious type. This device may cause serious injury or death if you continue to use it.
Affected Product
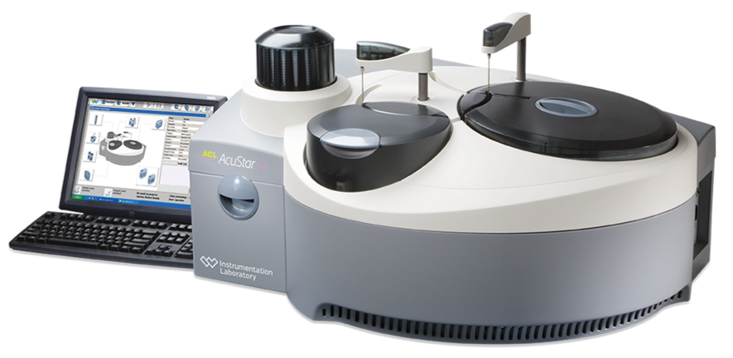
The FDA is aware that Instrumentation Laboratories (Werfen) has issued letters to affected customers recommending certain lots of HemosIL AcuStar HIT-IgG(PF4-H) be removed from where they are used or sold.
Affected devices:
| Product | Part No. | UDI | Lot No. | Expiration Date (YYYY-MM-DD) |
|---|---|---|---|---|
| HemosIL AcuStar HIT-IgG(PF4-H) | 0009800127 | 08426951126545 | B40005 | 2027-01-16 |
| HemosIL AcuStar HIT-IgG(PF4-H) | 0009800127 | 08426951126545 | B38062 | 2026-11-01 |
| HemosIL AcuStar HIT-IgG(PF4-H) | 0009802028 | 08426950545774 | B40006 | 2027-01-16 |
| HemosIL AcuStar HIT-IgG(PF4-H) | 0009802028 | 08426950545774 | B38063 | 2026-11-01 |
What to Do
Users should identify and destroy affected lots of HemosIL AcuStar HIT-IgG(PF4-H) according to the instructions below.
- Check your inventory for Lot No. B40005, B40006, B38062, and B38063 and destroy any remaining product following your facility’s standard procedures.
- Document the destruction on the Urgent Mandatory Response Tracking Form, including quantity destroyed and date of destruction.
- Identify an alternative method for of heparin-induced thrombocytopenia (HIT) antibody testing as a supply interruption is expected. Please work with your local Werfen representative for guidance and transition support.
Decisions regarding the review of previously reported patient results, patient follow-up, or repeat testing should be made by the laboratory and/or the treating clinician in accordance with local procedures, clinical judgement, and applicable regulatory requirements. For questions regarding specific patient results, please consult your laboratory medical director or other appropriate clinical leadership.
Reason for Recall
Instrumentation Laboratories (Werfen) stated that a performance issue has been identified affecting Lot No. B40005, B40006, B38062, and B38063 of HemosIL AcuStar HIT-IgG(PF4-H), Part No. 0009800127 and 0009802028. Customer complaints have reported obtaining an invalid calibration curve and in one complaint, a negative mean bias of 47% with patient samples. Werfen’s investigation has confirmed performance variability (negative bias of 20-35%) with the affected lots that may not be reliably detected through routine calibration and quality control testing. The root cause investigation is ongoing.
This performance issue may result in a negative bias that could cause sample results to be incorrectly reported below the assay cut-off (1.00 U/mL) and, therefore, be interpreted as “HIT IgG Antibody negative.” A false positive negative result may delay recognition of HIT and influence clinical management. In patients with HIT, continued exposure to heparin may increase the risk of thrombotic complications.
As of September 3, Instrumentation Laboratories (Werfen) has reported no serious injuries or deaths associated with this issue.
Device Use
The HemosIL AcuStar HIT-IgG(PF4-H) device is a qualitative, fully automated, chemiluminescent immunoassay (CIA) for the detection of IgG antibodies that react with the Platelet Factor 4 (PF4) when complexed to heparin. The result is interpreted as positive or negative and aids in determining the risk for HIT when used in conjunction with other laboratory and clinical findings.
Contact Information
Customers in the U.S. with adverse reactions, quality problems, or questions about this recall should contact Werfen Client Services at 1-800-955-9525.
Additional FDA Resources
Unique Device Identifier (UDI)
The unique device identifier (UDI) helps identify individual medical devices sold in the United States from distribution to use. The UDI allows for more accurate reporting, reviewing, and analyzing of adverse event reports so that devices can be identified more quickly, and as a result, problems potentially resolved more quickly.
How do I report a problem?
Health care professionals and consumers may report adverse reactions or quality problems they experienced using these devices to MedWatch: The FDA Safety Information and Adverse Event Reporting Program.